Why viscosity changes during chemical formulation development
Key Takeaways
- Used EV Battery Aftermarket Could Reach 10%–20% Share Within Five YearsAutomotive Tech Analyst
- Which electric steering components can cause intermittent assist loss?Automotive Tech Analyst
- Crusoe and Araico Plan 106 MW AI Cloud Infrastructure Project in ParaguayIT & Security Director
- Is an SDS enough to verify chemical compliance for polymers?Lead Materials Scientist
- Can Non-ISOBUS Implements Work With an ISOBUS Tractor? Retrofit OptionsAgri-Tech Strategist
- Titanium Dioxide Stability in Exterior Coatings: What Drives Chalking?Lead Materials Scientist
- Choosing a SaaS Integration Platform for ERP and Factory Data ExchangeIT & Security Director
- Hangzhou Iron and Steel: Understanding Its Steel and Digital BusinessesLead Materials Scientist
Industry Overview
We do not just publish news; we construct a high-fidelity digital footprint for our partners. By aligning with TNE, enterprises build the essential algorithmic "Trust Signals" required by modern search engines, ensuring they stand out to high-net-worth buyers in an increasingly crowded global digital landscape.
Why viscosity changes during chemical formulation development
A formulation that meets viscosity specifications in the lab can behave differently after scale-up, storage, or a change in raw material supplier. During chemical formulation development, these shifts often reflect changes in temperature, shear history, polymer interactions, pH, or ingredient compatibility—not simply measurement error. Identifying the mechanism is essential to distinguishing reversible processing effects from instability that could compromise product performance.
The practical question is not just “Why did the viscosity change?” It is “What changed in the material, and does that change matter during manufacturing, storage, or use?” Adding thickener before answering those questions can conceal a dispersion problem, reduce pumpability, or create a batch that passes release testing but fails later.
Before changing the formula, make the measurements comparable
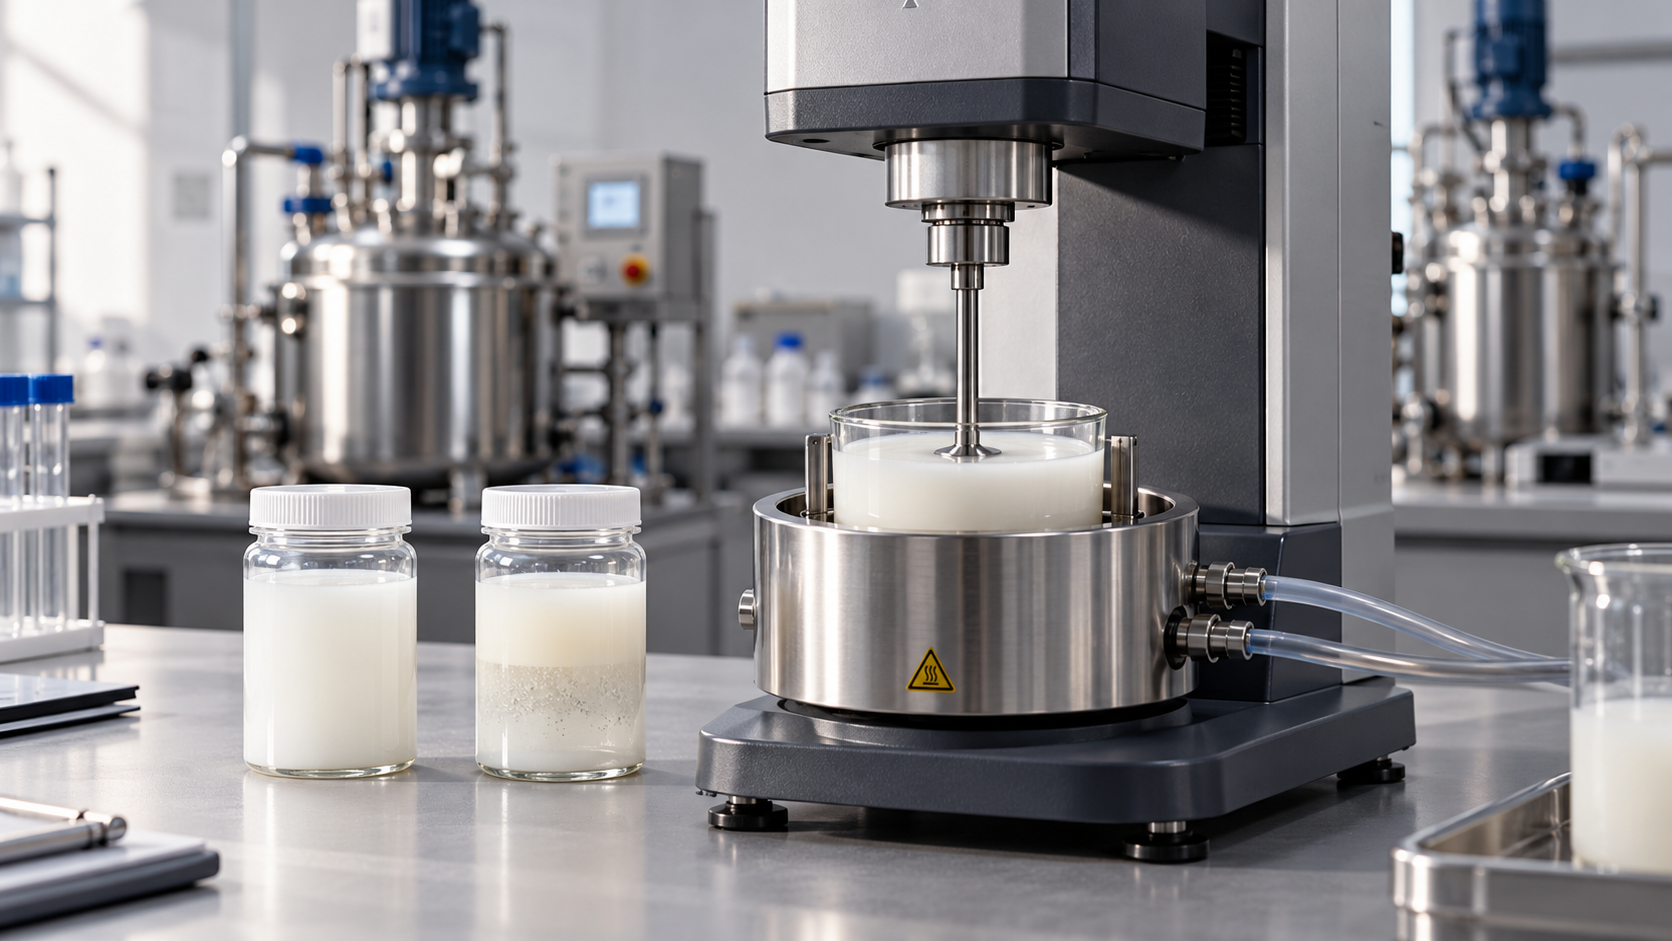
For many coatings, adhesives, cleaning products, and suspensions, viscosity is not a single material constant. These products are non-Newtonian: their apparent viscosity depends on the applied shear and sometimes on how long that shear acts. Two readings can both be correct while describing different material states.
A production sample measured warm immediately after mixing should not be compared directly with a laboratory sample measured after overnight rest. Likewise, matching rotational speed does not necessarily match shear conditions when instruments or measuring geometries differ. For some spindle arrangements, a unique shear rate cannot be assigned reliably.
A useful comparison records the instrument, spindle or geometry, speed or shear-rate program, actual sample temperature, sample age, loading procedure, pre-shear, rest period, and reading time. Confirm that torque is within the instrument’s recommended operating range. With structured suspensions, also consider wall slip, particle size relative to the measuring gap, and whether sampling disturbed the structure.
Entrained air deserves a separate check. Foaming can distort readings and make sample loading inconsistent. Deaeration may help, but aggressive vacuum can remove volatile ingredients. Use a preparation method that does not change the composition you intend to measure.
Temperature and shear can change viscosity without damaging the product
Many liquids become less viscous as temperature rises. In formulated systems, however, temperature also affects polymer solvation, surfactant organization, crystallization, and phase behavior. Some thermoresponsive systems thicken on heating; others lose structure abruptly. A generic temperature correction is therefore unsafe unless it has been validated for that formulation.
To investigate a warm batch that appears too thin, bring matched samples to the same temperature and allow adequate equilibration before retesting. If the difference disappears, the immediate issue may be the release-test procedure rather than the recipe. If it persists, examine whether heating changed the formulation irreversibly or accelerated another process.
Shear introduces another distinction. Shear thinning describes lower apparent viscosity at higher shear rates. Thixotropy describes time-dependent structural breakdown under shear and recovery afterward. A suspension may thin during pumping and rebuild in the filling tank. That can be desirable, provided recovery is fast enough to prevent settling or sagging.
Excessive processing can also cause lasting change. Certain high-molecular-weight polymers are vulnerable to mechanical degradation under severe conditions, while emulsions can experience changes in droplet size or stability. A low viscosity immediately after mixing does not prove damage; failure to recover under a controlled rest-and-retest procedure makes irreversible change more plausible.
A low-shear, high-shear, low-shear rheology sequence can help separate application thinning from recovery behavior. Its conditions should reflect the process being assessed, not simply an instrument’s default program.
The ingredients may still be building—or losing—their structure
Hydration and order of addition
A thickener is not necessarily fully functional when it first disappears into the liquid. Some polymers require time to wet, hydrate, dissolve, swell, or neutralize. Poor addition technique can create lumps with hydrated outer layers and dry interiors. The batch may initially look under-thickened, then gain viscosity as those regions slowly hydrate.
This is why addition order matters. Introducing a polymer into a concentrated salt solution may produce a different result from hydrating it in water before adding electrolytes. Other thickeners require specific activation or dispersion conditions. Follow the chemistry and supplier guidance for the grade being evaluated; there is no universal “thickener first” rule.
pH, salts, and associative interactions
For ionizable polymers, pH can change chain expansion and water interaction. Some alkali-swellable thickeners develop viscosity after neutralization, but the relationship is not indefinitely upward. Moving beyond a suitable operating window can reduce performance or destabilize other ingredients.
Electrolytes can screen polymer charges and lower thickening efficiency. Multivalent ions may interact more strongly, potentially promoting bridging or precipitation in susceptible systems. Water hardness, pigment slurries, neutralizers, and minor additives can all introduce ions that are absent from a simplified laboratory formula.
Associative thickeners add another complication: their performance depends partly on interactions with surfactants, binder particles, and other hydrophobic components. Changing a wetting agent or defoamer may alter viscosity even when thickener dosage stays constant. In surfactant-rich formulations, salt response can be non-monotonic—an addition may thicken the product up to a point, then thin it.
Dispersion, flocculation, and composition drift
A viscosity increase is not always evidence of successful thickening. Particle flocculation can create a weak network that raises low-shear viscosity while worsening appearance, flow, or storage behavior. Conversely, improved dispersion can lower viscosity by breaking particle networks. Interpret rheology alongside microscopy, dispersion checks, settling observations, or application testing as appropriate.
Composition can also drift. Water or solvent loss may concentrate a formulation; moisture uptake can change some reactive systems. Continued curing, hydrolysis, oxidation, or microbial activity may alter viscosity during storage. These possibilities require chemistry-specific investigation. A viscosity reading alone cannot identify which reaction occurred.
Why a successful laboratory batch can fail at production scale
Scaling a mixer by matching rpm is rarely sufficient. Vessel geometry, impeller design, circulation, power input, heat removal, and feed location all affect the material’s processing history. Matching tip speed alone does not guarantee equivalent mixing either. The relevant scale-up criterion depends on whether the limiting task is bulk blending, powder wetting, dispersion, or emulsification.
Consider a polymer powder added into a well-circulated laboratory vessel. At production scale, a faster addition rate into a poorly swept region can create local concentrations that prevent proper hydration. Extra mixing afterward may not fully correct the defect. The recipe is unchanged, but the route to the final structure is different.
Supplier changes need similar scrutiny. Materials sold under the same generic chemistry may differ in molecular-weight distribution, active content, particle size, surface treatment, or electrolyte content. Incoming viscosity specifications alone may not predict compatibility in the finished formulation.
For the materials and supply-chain questions covered by TradeNexus Edge (TNE), this distinction matters: commercial equivalence is not formulation equivalence. An alternate source should be qualified in the actual formulation and process, rather than accepted solely because its data sheet resembles the incumbent grade.
Investigate the pattern before choosing a correction
The direction and timing of a viscosity shift help narrow the investigation. They are clues, not diagnoses.
- Low immediately after mixing, then recovering: examine temperature, shear history, hydration time, and structural recovery before increasing thickener.
- Rising during storage: check delayed hydration, evaporation, flocculation, crystallization, and ongoing reaction. Compare container mass and appearance where relevant.
- Falling during storage: investigate pH drift, network disruption, degradation, and possible microbial activity where the chemistry permits it.
- Changing only after a supplier substitution: compare active content and relevant raw-material attributes, then test interactions in the full formulation.
- Different results between laboratories: exchange a retained sample and align conditioning and measurement procedures before reformulating.
Use controlled comparisons to test the strongest hypothesis. Retain a reference batch, change one variable at a time where practical, and document both processing conditions and sample history. If several factors are likely to interact, a small designed experiment is more informative than a sequence of unrelated recipe adjustments.
Set acceptance criteria around performance, not one number
A single-point viscosity specification can be useful for routine release once its relationship to performance is established. During chemical formulation development, it is often insufficient. Two products can share the same measured viscosity yet behave differently during pumping, spraying, leveling, dispensing, or storage.
Choose tests around the failure mode. A suspension may need low-shear structure and settling assessments; a spray coating may require application-shear behavior and post-spray recovery checks. A dispensable adhesive may need evaluation after realistic pumping and rest cycles. Yield stress can be useful, but its measured value depends on the method and should not be treated as an absolute guarantee against settling.
Storage studies should use relevant packaging, temperatures, and sampling intervals. Accelerated exposure can reveal weaknesses, but it may trigger mechanisms that do not occur under normal storage and does not automatically establish shelf life.
Before production approval, establish which viscosity changes are reversible, which indicate deterioration, and which affect the customer’s process. The next development batch should test that distinction—not merely bring the release reading back inside its limits.
Deep Dive
Related Intelligence
 Specialty ChemicalsSodium Methoxide Selection: How Chemical Buyers Reduce Specification and Handling RiskAnalyst:Lead Materials Scientist
 Specialty ChemicalsHow Corporate Case Studies Reveal Real-World Challenges in Adopting Specialty Chemicals for Food SafetyAnalyst:Lead Materials Scientist
 Specialty ChemicalsCan Brand Elevation Tactics Reduce Customer Acquisition Cost for Specialty Chemicals in Organic Food Production?Analyst:Lead Materials Scientist